

| Size | Price | Stock | Qty |
|---|---|---|---|
| 50mg |
|
||
| 100mg |
|
||
| 1g |
|
||
| 2g |
|
||
| 5g |
|
||
| Other Sizes |


Purity: ≥98%
| Targets |
5-HT3 Receptor
Ondansetron HCl (GR 38032; SN 307; GR-C507/75) is a highly selective antagonist of the 5-hydroxytryptamine 3 (5-HT₃) receptor. In human recombinant 5-HT₃ receptors (expressed in HEK 293 cells), it exhibits a Ki value of 0.16 nM; in rat enterochromaffin cell membranes, the Ki for 5-HT₃ receptors is 0.3 nM [2] - Ondansetron HCl (GR 38032; SN 307; GR-C507/75) has negligible affinity for other 5-HT receptor subtypes (5-HT₁A, 5-HT₂A: Ki > 10,000 nM) and neurotransmitter receptors (dopamine D₂, muscarinic M₁: Ki > 5000 nM) in human brain membranes [2] |
|---|---|
| ln Vitro |
In vitro activity: Ondansetron produces no change in salivation, rectal temperature, or urination, but it lessens the intensity of withdrawal signs like increased defecation, jumping, and wet-dog shakes. It also elevates the nociceptive threshold values, which are decreased by precipitated withdrawal. Rats' gastric emptying of glass beads is markedly improved by ondansetron and granisetron, while the cisplatin-induced slowing of gastric emptying is ameliorated. In the forced swim and tail suspension tests, ondansetron shows a biphasic dose-response profile in mice, with antidepressant-like effects peaking at 0.1 mg/kg. Desipramine and 8-hydroxy-2-(di-n-propylamino) tetralin do not affect the effects of ondansetron pretreatment, but it does increase the antidepressant effects of fluoxetine and venlafaxine. In the open field, ondansetron (10 mg/kg) reverses hyperactivity. In the elevated plus maze, it also reduces the percentage of entries and the amount of time spent in open arms. At a dose of 0.01 mg/kg, but not at 0.1 mg/kg, ondansetron, a potent and selective 5HT3 receptor antagonist, is demonstrated to be effective in preventing the disruption of LI caused by amphetamine. Ondansetron exhibits the ability to reduce spikes in dopamine activity and works pharmacologically with amphetamine without changing baseline levels of dopamine activity. Ondansetron improves behavior in both young adults and older animals. It also prevents scopolamine, electrolesions, and ibotenic acid lesions of the nucleus basalis magnocellularis from impairing habituation. A scopolamine-induced impairment is counteracted by ondansetron and arecoline.
5-HT₃ Receptor Antagonism (Calcium Mobilization): In NG108-15 cells (expressing native 5-HT₃ receptors), Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (10⁻¹¹ to 10⁻⁶ M) concentration-dependently blocks 5-HT (1 μM)-induced intracellular calcium elevation: 10⁻⁹ M inhibits the calcium response by 50% (IC₅₀ = 0.8 nM), with complete inhibition at 10⁻⁷ M [2] - Receptor Selectivity Assay: In human brain membrane preparations, Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (1 μM) shows no significant displacement of [³H]-spiperone (D₂ ligand) or [³H]-pirenzepine (M₁ ligand), confirming lack of off-target binding to non-5-HT₃ receptors [2] - Enterochromaffin Cell 5-HT Release: In isolated rat intestinal enterochromaffin cells, Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (100 nM, 1 μM) has no effect on basal or depolarization-induced 5-HT release, indicating it does not interfere with 5-HT synthesis or secretion [2] |
| ln Vivo |
prevents radiation sickness when combined with CP-99,994 (15 mg/kg) and Dexamethasone (2 mg/kg), in radiation-induced pica model[1].
Mouse Radiation Sickness Model: In male ICR mice exposed to 8 Gy whole-body γ-radiation, intraperitoneal (i.p.) administration of Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (1, 3, 10 mg/kg) 30 min pre-radiation dose-dependently reduces radiation-induced symptoms: 10 mg/kg decreases vomiting episodes by 80% (vs. 100% in vehicle), prevents weight loss (maintains 90% baseline weight at day 3 vs. 70% in vehicle), and increases 7-day survival rate from 40% (vehicle) to 75% [1] - Rat Cisplatin-Induced Emesis Model: In male Sprague-Dawley rats treated with cisplatin (6 mg/kg, i.p., chemotherapeutic), oral Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (0.3, 1, 3 mg/kg) 1 h pre-cisplatin dose-dependently inhibits acute emesis: 3 mg/kg reduces emetic episodes by 90% (vs. 60 episodes in vehicle) and delays emesis onset from 2 h to 6 h [2] - Ferret Post-Operative Nausea Model: In female ferrets undergoing laparotomy, intravenous (i.v.) Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (0.1, 0.3 mg/kg) 10 min pre-surgery reduces post-operative nausea (measured via behavioral signs: retching, pawing at mouth) by 70% (0.3 mg/kg) vs. vehicle [2] |
| Enzyme Assay |
Rat Intestinal Membrane 5-HT₃ Binding Assay: Rat proximal intestine was dissected, homogenized in ice-cold Tris-HCl buffer (50 mM, pH 7.4, containing 120 mM NaCl, 5 mM KCl) and centrifuged at 48,000 × g for 15 min. The membrane pellet was resuspended in the same buffer. 50 μg of membrane protein was incubated with [³H]-GR65630 (0.5 nM, a selective 5-HT₃ ligand) and various concentrations of Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (10⁻¹² to 10⁻⁶ M) at 25°C for 60 min. Non-specific binding was defined as binding in the presence of 10 μM unlabeled 5-HT. Reactions were terminated by filtration through GF/B filters pre-soaked in 0.1% polyethyleneimine, washed 3 times with ice-cold buffer, and radioactivity counted via liquid scintillation spectrometry. Ki values were calculated using the Cheng-Prusoff equation [2]
|
| Cell Assay |
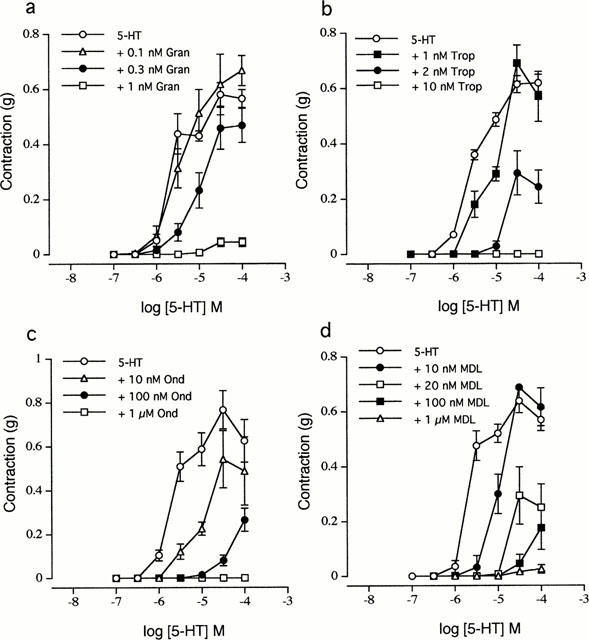
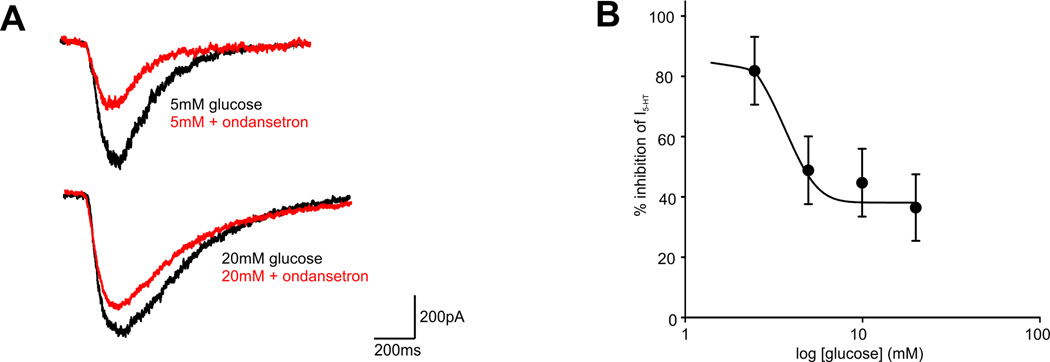
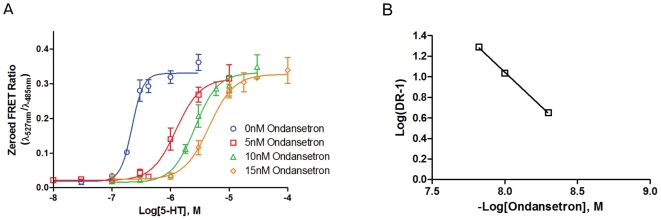
Ondansetron, an antagonist of the 5-HT3 receptor, inhibited the transient inward currents that 5-HT evoked (EC50 = 3.4 microM; Hill coefficient = 1.8) (IC50 = 103 pM). Ondansetron, a 5-HT3A receptor antagonist, reversibly inhibited the 5-HT (30 microM) signal by 70% at 0.3 nM and completely eliminated the response at 3 nM.
Cell Culture & Treatment: NG108-15 cells (mouse neuroblastoma-rat glioma hybrid) were cultured in DMEM/Ham’s F12 medium supplemented with 10% fetal bovine serum (FBS), 100 U/mL penicillin, and 100 μg/mL streptomycin at 37°C in 5% CO₂. Cells were seeded in 96-well black-walled plates (1×10⁴ cells/well) and allowed to adhere for 24 h [2] - Calcium Detection: Medium was replaced with HEPES-buffered saline (HBS: 140 mM NaCl, 5 mM KCl, 1 mM CaCl₂, 10 mM HEPES, pH 7.4) containing the fluorescent calcium indicator Fluo-4 AM (4 μM) and Pluronic F-127 (0.02%) for 45 min at 37°C. Cells were washed with HBS, then treated with Ondansetron HCl (GR 38032; SN 307; GR-C507/75) (10⁻¹¹ to 10⁻⁶ M) for 10 min, followed by 5-HT (1 μM). Fluorescence intensity (excitation 488 nm, emission 525 nm) was measured every 2 s for 5 min using a microplate reader [2] |
| Animal Protocol |
0.1 mg/kg, 0.33 and 1 mg/kg, i.p.
Mice Mouse Radiation Sickness Model: Male ICR mice (25–30 g) were housed under 12 h light/dark cycles with free access to food and water. Mice were randomized into 4 groups (n=10/group): Vehicle (normal saline + 0.1% DMSO, i.p.), Ondansetron HCl 1 mg/kg (i.p.), 3 mg/kg (i.p.), 10 mg/kg (i.p.). Ondansetron HCl (GR 38032; SN 307; GR-C507/75) was dissolved in normal saline with 0.1% DMSO (injection volume: 10 mL/kg). Thirty minutes after drug administration, mice were exposed to 8 Gy γ-radiation (dose rate: 1 Gy/min). Vomiting episodes were counted every 12 h for 3 days; body weight was measured daily; survival was recorded for 7 days [1] - Rat Cisplatin-Induced Emesis Model: Male Sprague-Dawley rats (250–280 g) were fasted for 12 h (water ad libitum) and randomized into 4 groups (n=8/group): Vehicle (0.5% methylcellulose, p.o.), Ondansetron HCl 0.3 mg/kg (p.o.), 1 mg/kg (p.o.), 3 mg/kg (p.o.). One hour after oral gavage, rats received cisplatin (6 mg/kg, i.p.) dissolved in normal saline. Emetic episodes (defined as oral expulsion of gastric contents) were counted every 30 min for 24 h; emesis onset time was recorded [2] |
| ADME/Pharmacokinetics |
Oral absorption: In healthy volunteers (n=6), oral administration of ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) (8 mg) reached a peak plasma concentration (Cmax) of 24 ng/mL at 1.5 hours (Tmax), with an absolute oral bioavailability of 60% (partially metabolized by the liver) [2]
- Metabolism: Ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) is mainly metabolized in the liver by cytochrome P450 enzymes CYP3A4 (major) and CYP2D6 (minor), forming inactive metabolites (e.g., 8-hydroxyondansetron). No significant metabolism of CYP1A2 or CYP2C9 was observed [2] - Excretion and half-life: In humans, the terminal elimination half-life (t₁/₂) of ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) is 3.5 hours. Approximately 70% of the administered dose is excreted in the urine (as metabolites) within 72 hours, and 20% in the feces; <5% is excreted unchanged[2] - Tissue distribution: In male rats, oral administration of ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) (1 mg/kg) resulted in high gastrointestinal tissue concentrations: 1 hour after administration, the intestinal mucosal concentration was 180 ng/g, and the plasma concentration was 22 ng/g (tissue/plasma ratio = 8.2), with low central nervous system penetration (brain concentration = 5 ng/g)[2] |
| Toxicity/Toxicokinetics |
Effects During Pregnancy and Lactation
◉ Overview of Use During Lactation Ondansetron is commonly used to relieve nausea during and after cesarean section, usually at an intravenous dose of 4 to 8 mg. Use of ondansetron during and after cesarean section does not appear to affect the initiation of breastfeeding. No adverse reactions have been reported in infants of women who received ondansetron postpartum, nor in pharmacokinetic studies. Studies on the use of ondansetron in breastfeeding women outside the early postpartum period are insufficient, but the drug is approved for use in infants up to 1 month old. Computer models show that drug concentrations in breast milk are much lower than this dose. No special precautions are required. ◉ Effects on Breastfed Infants In a pharmacokinetic study of 78 women who received intravenous ondansetron postpartum, no adverse reactions were reported in their breastfed infants. ◉ Effects on Lactation and Breast Milk A randomized, double-blind study compared the effects of placebo versus intravenous 4 mg ondansetron after cesarean section in preventing postoperative nausea and vomiting. There was no difference in the time to first breastfeeding between the two groups. In a retrospective study of women undergoing cesarean section, three regimens were compared: dexmedetomidine before anesthesia and during delivery (n = 115), saline before anesthesia and during delivery, and dexmedetomidine postpartum (n = 109), and saline before anesthesia and during delivery (n = 168). All women received 4 mg ondansetron as needed, prior to suture removal. The mean total ondansetron intake ranged from 6 mg to 9 mg across all groups. The time to first lactation was similar across all groups (25 to 28 minutes). Plasma protein binding: In human plasma (as determined by ultrafiltration), the protein binding of ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) was 73% in the concentration range of 10–1000 ng/mL, and was concentration-independent [2] - Acute toxicity: In male ICR mice, the oral LD₅₀ of ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) was >2000 mg/kg; and in rats, the intraperitoneal LD₅₀ was >1000 mg/kg. In rats, no death or serious toxicity (convulsions, respiratory depression) was observed at doses up to 500 mg/kg [2] - Clinical adverse reactions: In a phase III clinical trial (n=1500 patients with chemotherapy-induced nausea/vomiting), common adverse reactions to ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) (8 mg, twice daily) included headache (15%), constipation (10%), and dizziness (5%); these adverse reactions were mild to moderate and resolved without dose adjustment [2] - Drug interactions: In humans, co-administration of ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) (8 mg, orally) with ketoconazole (a CYP3A4 inhibitor, 400 mg/day) increased the Cmax of ondansetron by 1.8-fold and prolonged t₁/₂ to 5.2. The time is short, but it does not increase adverse reactions [2] |
| References | |
| Additional Infomation |
Ondansetron hydrochloride is the hydrochloride salt of the racemic mixture of ondansetron, a carbazole derivative and a selective competitive 5-HT3 receptor antagonist with antiemetic activity. Although its mechanism of action is not fully elucidated, ondansetron appears to competitively block the action of 5-HT3 receptors in the peripheral gastrointestinal tract and the postmedulatal medulla oblongata (the region containing the chemoreceptor trigger zone (CTZ) of the central nervous system), thereby inhibiting nausea and vomiting induced by chemotherapy and radiotherapy. It is effective in treating nausea and vomiting induced by cytotoxic chemotherapy drugs (including cisplatin) and has anti-anxiety and antipsychotic effects.
See also: Ondansetron (containing the active ingredient). Drug Indications Treatment of alcohol use disorder Ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) is a first-generation 5-HT₃ receptor antagonist that was approved by the FDA in 1991 for the prevention of chemotherapy-induced nausea and vomiting (CINV), radiation-induced nausea and vomiting (RINV), and postoperative nausea and vomiting (PONV) [2]. - Mechanism of action: Its antiemetic effect is achieved by blocking 5-HT₃ receptors in the gastrointestinal tract (peripheral) and the nervous system (nervous system). Chemoreceptor trigger zone (CTZ, central nervous system): 5-HT released from damaged intestinal cells activates 5-HT₃ receptors, triggering the vomiting reflex; ondansetron inhibits this activation [1,2] - New application (radiation sickness): Ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) (10 mg/kg, intraperitoneal injection) in combination with dexamethasone (2 mg/kg) and an NK1 receptor antagonist completely blocked radiation-induced vomiting in mice, supporting its potential for treating radiation sickness in humans [1] - Dosage advantage: Unlike older generation antiemetics (e.g., metoclopramide), ondansetron hydrochloride (GR 38032; SN 307; GR-C507/75) has no extrapyramidal side effects, making it more suitable for long-term use in cancer patients [2] |
| Molecular Formula |
C18H19N3O
|
|---|---|
| Molecular Weight |
329.82
|
| Exact Mass |
329.129
|
| Elemental Analysis |
C, 65.55; H, 6.11; Cl, 10.75; N, 12.74; O, 4.85
|
| CAS # |
99614-01-4
|
| Related CAS # |
Ondansetron hydrochloride dihydrate; 103639-04-9; Ondansetron; 99614-02-5; Ondansetron-d3 hydrochloride; 1346605-02-4; Ondansetron-d6 hydrochloride; 1225442-22-7
|
| PubChem CID |
68647
|
| Appearance |
Solid powder
|
| Density |
1.27g/cm3
|
| Boiling Point |
546ºC at 760mmHg
|
| Melting Point |
178.5-179.5ºC
|
| Flash Point |
284ºC
|
| LogP |
3.93
|
| Hydrogen Bond Donor Count |
1
|
| Hydrogen Bond Acceptor Count |
2
|
| Rotatable Bond Count |
2
|
| Heavy Atom Count |
23
|
| Complexity |
440
|
| Defined Atom Stereocenter Count |
0
|
| SMILES |
O=C1C(CN2C=CN=C2C)CCC(N3C)=C1C4=C3C=CC=C4.[H]Cl
|
| InChi Key |
MKBLHFILKIKSQM-UHFFFAOYSA-N
|
| InChi Code |
InChI=1S/C18H19N3O.ClH/c1-12-19-9-10-21(12)11-13-7-8-16-17(18(13)22)14-5-3-4-6-15(14)20(16)2;/h3-6,9-10,13H,7-8,11H2,1-2H3;1H
|
| Chemical Name |
9-methyl-3-[(2-methylimidazol-1-yl)methyl]-2,3-dihydro-1H-carbazol-4-one;hydrochloride
|
| Synonyms |
GR 38032F; GRC 50775; SN 307; GR-38032F; GRC-50775; SN-307; GR38032F; GRC50775; SN307; GR 38032F; trade name: Zofran
|
| HS Tariff Code |
2934.99.9001
|
| Storage |
Powder -20°C 3 years 4°C 2 years In solvent -80°C 6 months -20°C 1 month Note: (1). Please store this product in a sealed and protected environment (e.g. under nitrogen), avoid exposure to moisture and light. (2). This product is not stable in solution, please use freshly prepared working solution for optimal results. |
| Shipping Condition |
Room temperature (This product is stable at ambient temperature for a few days during ordinary shipping and time spent in Customs)
|
| Solubility (In Vitro) |
|
|||
|---|---|---|---|---|
| Solubility (In Vivo) |
Solubility in Formulation 1: ≥ 5 mg/mL (15.16 mM) (saturation unknown) in 10% DMSO + 40% PEG300 + 5% Tween-80 + 45% Saline (add these co-solvents sequentially from left to right, and one by one), Clear solution.
For example, if 1 mL of working solution is to be prepared, you can add 100 μL of DMSO stock solution (50.0 mg/mL) to 400 μL of PEG300 and mix well; then add 50 μL of Tween-80 and mix well; finally add 450 μL of physiological saline and adjust the volume to 1 mL. Preparation of saline: Dissolve 0.9 g of sodium chloride in 100 mL ddH₂ O to obtain a clear solution. Solubility in Formulation 2: ≥ 5 mg/mL (15.16 mM) (saturation unknown) in 10% DMSO + 90% (20% SBE-β-CD in Saline) (add these co-solvents sequentially from left to right, and one by one), Clear solution. For example, if 1 mL of working solution is to be prepared, you can add 100 μL of DMSO stock solution (50.0 mg/mL) to 900 μL of 20% SBE-β-CD saline and mix well. Preparation of 20% SBE-β-CD in Saline (4°C,1 week): Dissolve 2 g SBE-β-CD in 10 mL saline to obtain a clear solution. Solubility in Formulation 3: ≥ 5 mg/mL (15.16 mM) (saturation unknown) in 10% DMSO + 90% Corn Oil (add these co-solvents sequentially from left to right, and one by one), Clear solution. For example, if 1 mL of working solution is to be prepared, you can add 100 μL of DMSO stock solution (50.0 mg/mL) to 900 μL of corn oil and mix well. (Please use freshly prepared in vivo formulations for optimal results.) |
| Preparing Stock Solutions | 1 mg | 5 mg | 10 mg | |
| 1 mM | 3.0320 mL | 15.1598 mL | 30.3196 mL | |
| 5 mM | 0.6064 mL | 3.0320 mL | 6.0639 mL | |
| 10 mM | 0.3032 mL | 1.5160 mL | 3.0320 mL |
*Note: Please select an appropriate solvent for the preparation of stock solution based on your experiment needs. For most products, DMSO can be used for preparing stock solutions (e.g. 5 mM, 10 mM, or 20 mM concentration); some products with high aqueous solubility may be dissolved in water directly. Solubility information is available at the above Solubility Data section. Once the stock solution is prepared, aliquot it to routine usage volumes and store at -20°C or -80°C. Avoid repeated freeze and thaw cycles.
Calculation results
Working concentration: mg/mL;
Method for preparing DMSO stock solution: mg drug pre-dissolved in μL DMSO (stock solution concentration mg/mL). Please contact us first if the concentration exceeds the DMSO solubility of the batch of drug.
Method for preparing in vivo formulation::Take μL DMSO stock solution, next add μL PEG300, mix and clarify, next addμL Tween 80, mix and clarify, next add μL ddH2O,mix and clarify.
(1) Please be sure that the solution is clear before the addition of next solvent. Dissolution methods like vortex, ultrasound or warming and heat may be used to aid dissolving.
(2) Be sure to add the solvent(s) in order.
| NCT Number | Recruitment | interventions | Conditions | Sponsor/Collaborators | Start Date | Phases |
| NCT04169828 | Active Recruiting |
Drug: Methotrexate Drug: Ondansetron |
Juvenile Idiopathic Arthritis | University of British Columbia | August 2, 2019 | Not Applicable |
| NCT04209595 | Active Recruiting |
Drug: PLX038 Drug: Ondansetron |
Small Cell Lung Cancer Extra-Pulmonary Small Cell Carcinomas |
National Cancer Institute (NCI) |
April 8, 2020 | Phase 1 Phase 2 |
| NCT05378113 | Recruiting | Drug: Ondansetron Drug: Lidocaine |
Injection Site Irritation | Emory University | May 18, 2023 | Phase 2 |
| NCT05620641 | Recruiting | Drug: Ondansetron 8mg Drug: Gabapentin |
Sleeve Gastrectomy Morbid Obesity |
Tanta University | October 1, 2022 | Phase 3 |
| NCT03865290 | Recruiting | Drug: Ondansetron 8mg Drug: Placebo |
Indigestion Diabetes Mellitus |
Mayo Clinic | April 2, 2019 | Phase 2 |
 |
|---|
 |
 |
Products are for research use only; We do not sell to patients
Copyright 2020 InvivoChem LLC | All Rights Reserved


 COA