

| Size | Price | Stock | Qty |
|---|---|---|---|
| 5mg |
|
||
| 10mg |
|
||
| 25mg |
|
||
| 50mg |
|
||
| 100mg | |||
| 250mg | |||
| Other Sizes |
Purity: ≥98%
SB-334867 HCl (SB-334867A; SB334867; SB 334867), the hydrochloride salt of SB-334867, is a novel, non-peptide, selective orexin-1 (OX1) receptor antagonist with the potential for treating sleeping disorders. It inhibits orexin-1 with a pKb value of 7.2. It was the first non-peptide antagonist to be developed with selectivity for the orexin receptor subtype OX1, with around 50x selectivity for OX1 over OX2 receptors. It has been demonstrated to have sedative and anorectic effects in animals. It has also proven helpful in defining how orexinergic regulation of brain systems related to appetite, sleep, and other physiological processes is regulated.
| Targets |
OX1 receptor
|
|---|---|
| ln Vitro |
In vitro activity: SB-334867 has no effect on the UTP (3 microM)-induced calcium response, but it inhibits the orexin-A (10 nM) and orexin-B (100 nM)-induced calcium responses in CHO-OX1 cells, with pKB values of 7.27 and 7.23, respectively.
|
| ln Vivo |
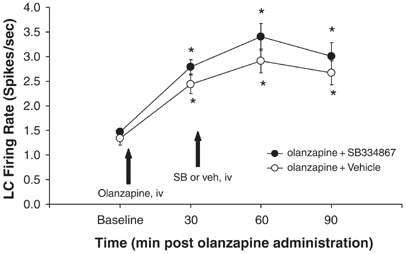
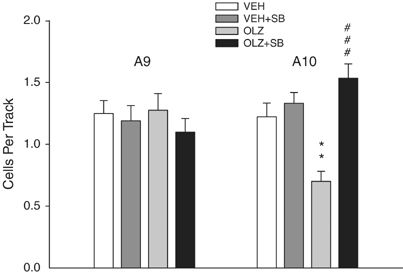
SB-334867 (30 mg/kg, i.p.) dramatically lowers food intake in both naturally occurring and orexin-A-induced rats, both male and female. In rats, the effects of antipsychotic medications on dopamine neuronal activity are blocked by SB-334867 (2 mg/kg, i.v.). Additionally, SB-334867 prevents rats from developing a tolerance to morphine analgesics.
|
| Enzyme Assay |
SB-334867 free base has an IC50 value of 7.2 (pKb) and is a selective non-peptide orexin OX1 receptor antagonist. In CHO-OX(1) cells, SB-334867-A did not affect the calcium response induced by UTP (3 microM) but it did suppress the responses to orexin-A (10 nM) and orexin-B (100 nM) (pK(B)=7.27+/-0.04 and 7.23+/-0.03, respectively, n=8). The calcium responses mediated by OX(2) were also inhibited by SB-334867-A (10 microM) (32.7+/-1.9% versus orexin-A).
|
| Cell Assay |
Two peptides extracted from the rat hypothalamus are called orexin-A and orexin-B. Some physiological processes that they are involved in include feeding regulation, energy metabolism, and sleep-wake cycle regulation. In CHO-OX1 cells, SB-334867 can suppress the calcium responses induced by orexin-A and orexin-B, with pKB values of 7.27 and 7.23, respectively. For OX1 receptors, SB-334867 exhibits greater selectivity than OX2 receptors. It inhibits the calcium responses in CHO-OX2 cells that are induced by orexin-A by 32.7% and orexin-B by 22%, respectively.
|
| Animal Protocol |
Dissolved in 10% (w/v) Encapsin in sterile water; 30 mg/kg; i.p. administration
Male and female Sprague–Dawley rats Drugs in Behavioral Experiments[1] The following drugs were used in the experiments: morphine hydrochloride trihydrate (Cosmetic Pharma, Poland) and 1-(2-methyylbenzoxanzol-6-yl)-3-[1,5]naphthyridin-4-yl-urea hydrochloride (SB-334867)—a selective OX-1 receptor antagonist. Morphine was dissolved in 0.9% saline, and SB-334867 was dissolved in three drops of DMSO and diluted in 0.9% saline (final the DMSO concentration 0.1%). All used substances were delivered intraperitoneally (i.p.) in a volume of 10 ml/kg. Morphine was used at the dose of 10 mg/kg, and SB-334867 was injected at the dose of 20 mg/kg. As literature data show, the minimal effective dose of SB-334867 is 30 mg/kg. According to the generally accepted principles of behavioral sensitization research, an ineffective dose of pharmacological agents (SB-334867 in our study) is recommended. Therefore, based on the literature data and on our preliminary unpublished results, the dose of SB-334867 (20 mg/kg), administered in the reported study, was subthreshold. The animals in a control group received the same volume of saline at the respective time point before the test. Procedure of Behavioral Sensitization[1] The Influence of SB-334867(OX-1 Receptor Antagonist, 20 mg/kg, i.p.) on the Acquisition of Morphine Sensitization to Locomotor Activity[1] The induced morphine behavioral sensitization in mice was based on the method, described by Kuribara, with a modification of Kotlińska and Bocheński. The animals received five injections of morphine (i.p.) at the dose of 10 mg/kg every 3 days (on the 1st, 4th, 7th, 10th, and 13th day of the experiment). Seven days after the last morphine injection (on the 20th day of the study), the mice were administered with a challenge dose of morphine (10 mg/kg, i.p.). Aiming to grade the development of behavioral sensitization, the mice were immediately placed into the actometer to record their locomotor activity for the period of 60 min. The control animals were administered with saline (i.p.).[1] Afterwards, the effects of SB-334867 (the selective OX-1 receptor antagonist) on the acquisition of morphine-induced sensitization were explored. SB-334867 was administered 15 min before morphine injection on the 1st, 4th, 7th, 10th, and 13th day of the experiment, but not on the 20th day. The control animals were administered with saline (i.p.). All 32 P rats had chronic access to ethanol in the home cage for approximately 8–14 months before the current studies were conducted. P rats were divided into 3 groups. One group (n = 10) was used to test the effects of SB-334867 and a second group (n = 11) was used to test the effects of LSN2424100 (one rat was excluded from the experiment due to low baseline drinking). A within-subjects experimental design was used to test the OX1 and OX2 receptor antagonists. These rats, along with another group of 11 (i.e., all 32 P rats) were tested in the almorexant study using a between-subjects design (n = 8/dose).[2] N-((1H-imidazol-2-yl)methyl)-N-([1,1′-biphenyl]-2-yl)-4-fluorobenzenesulfonamide hydrochloride (LSN2424100), SB-334867, (S)-almorexant (ACT-078573), and the inactive (R) enantiomer of almorexant were synthesized at Lilly Research Laboratories (Indianapolis, IN). Naltrexone hydrochloride was purchased from Sigma Aldrich (St. Louis, MO). For rat experiments, the OX1 antagonist SB-334867 was dissolved in a vehicle of 10% (2-hydroxypropyl)-β-cyclodextrin, 2% dimethyl sulfoxide, and 0.05% lactic acid in water, and administered by intraperitoneal (i.p.) injection in a dose volume of 1 ml/kg. The OX2 antagonist LSN2424100 was suspended in 1% carboxymethyl cellulose, 0.25% polysorbate-80 and 0.05% Dow antifoam in water, and administered by i.p. injection in a dose volume of 1 ml/kg. The mixed OX1/2 antagonist almorexant, and its inactive enantiomer, were dissolved in a 20% Captisol solution and administered orally (p.o.) in a dose volume of 1 ml/kg. Naltrexone was dissolved in water with the addition of 15 μl 85% lactic acid.[2] For mouse experiments, SB-334867 was dissolved using 0.01% polysorbate-80 in saline. Almorexant was dissolved in 20% Captisol in water. LSN2424100 was suspended using 1% carboxymethyl cellulose and 0.25% polysorbate-80 in water. All compounds were administered by i.p. injection at a dose volume of 10 ml/kg. Home cage 2-bottle choice drinking in P rats[2] P rats were housed individually in TSE LabMaster cages with food, water, and 15% ethanol (v/v) available at all times. Water and ethanol intake (in ml) were measured once every 5 min throughout the 12-h dark cycle and recorded for later analysis. In the first experiment, rats (n = 10) received vehicle, 3, 10, or 30 mg/kg SB-334867 (i.p.), 60 min before onset of the 12-h dark phase of the light-dark cycle, using a within-subjects design. In the second experiment, rats (n = 10) received vehicle, naltrexone (10 mg/kg), or LSN2424100 at doses of 10 or 30 mg/kg (i.p.), 60 min before onset of the 12-h dark phase, using a within-subjects design (one rat was excluded from the experiment due to low baseline drinking). In the third experiment, rats (n = 32) received vehicle, naltrexone (10 mg/kg), or S-almorexant at doses of 60 or 100 mg/kg (p.o.), 60 min before onset of the dark cycle, using a between-subjects design. Naltrexone was included in the study design as a positive control, since this dose of naltrexone has been shown to effectively reduce ethanol consumption in P rats under these testing conditions. For all experiments, a 60-min pre-treatment period was chosen so that the onset of the dark cycle roughly coincided with the time at which maximal brain concentrations were achieved (data not reported). Consumption of water and ethanol was measured during the first 3 h of the dark cycle, based on the short half-lives and high metabolism of the compounds. Experiments were conducted using a within-subject design, with 3–4 days washout between administration of different doses, which were counterbalanced using a Latin square design. One group of n = 10 rats was used to test the effects of SB-334867, LSN2424100, and almorexant on operant responding maintained on a progressive ratio schedule, in separate experiments. Drugs were administered two days per week (Tues and Fri) to allow for washout between subsequent doses. Rats received vehicle, 3, 10, or 30 mg/kgSB-334867 (i.p., 30 min prior to the session); vehicle, 3, 10, or 30 mg/kg LSN2424100 (i.p., 30 min prior to the session); or vehicle, 10, 30, or 60 mg/kg almorexant or 60 mg/kg of the inactive enantiomer of almorexant (p.o., 60 min prior to the session). On all other days, rats received progressive ratio operant testing without any drug treatments to maintain operant performance and confirm return to baseline behaviors. One rat was excluded from testing 60 mg/kg almorexant due to observation of a skin rash not related to the study drug.[2] Binge drinking in C57BL/6J mice[2] One week prior to ethanol intake testing, mice were given daily saline injections (i.p.) to acclimate them to handling and injection procedures. Ethanol consumption was assessed using a 4-day drinking-in-the-dark (DID) paradigm during which the water bottle in the home cage was replaced with a single bottle of ethanol (20% v/v) starting 3 h after the onset of the dark cycle. This procedure has been shown to produce high blood ethanol concentrations (BECs) resulting from high levels of ethanol consumption in a relatively short period of time (Rhodes et al., 2005). On the first three days, animals were injected with saline or vehicle 30 min prior to a 2-h period of access to ethanol. On the 4th day, drugs were administered via i.p. injection 30 min prior to the test session, which was extended to 4 h. One cohort of mice was administered vehicle, 3, 10, or 30 mg/kg SB-334867 (n = 10/dose). A second cohort of mice was tested with vehicle, 15, 30, or 60 mg/kg LSN2424100 (n = 9–10/dose). A third cohort of animals was given vehicle, 25, 50, or 100 mg/kg almorexant (n = 10/dose). In order to assess resulting BECs, immediately upon removal of ethanol bottles, blood samples were collected from the retro-orbital sinus and centrifuged. [2] In order to assess the specificity of drug effects on ethanol consumption, an additional group of ethanol-naïve animals was tested with sucrose solution (1% w/v) in the same DID paradigm (Days 1–3: 2-h access with saline injections; Day 4: 4-h test session with drug pretreatment). On the 4th day, vehicle, 3, 10, or 30 mg/kg (n = 6–7/dose) SB-334867 was administered prior to the 4-h access period. During a subsequent week of testing, these same mice were administered either vehicle or 100 mg/kg almorexant (n = 14/dose) before the 4-h intake session. In a separate cohort of mice, vehicle or 60 mg/kg LSN2424100 (n = 9–10/dose) was administered prior to the 4-h test. |
| References |
|
| Additional Infomation |
1-(2-Methyl-1,3-benzoxazol-6-yl)-3-(1,5-naphthid-4-yl)urea is a naphthidine derivative. This study focuses on the role of the orexin-1 receptor antagonist SB-334867 in morphine-induced motor sensitization in mice. Behavioral sensitization refers to an enhanced systemic response to the same dose of an addictive substance, which is thought to increase drug cravings and the risk of relapse. Morphine sensitization was induced in mice by intermittent injections of morphine (10 mg/kg, intraperitoneally, every 3 days for a total of 5 injections), followed by a challenge dose of morphine (10 mg/kg) 7 days later. To assess the effect of orexin system blockade on the acquisition of sensitization, SB-334867 was administered before each morphine injection, except for the challenge dose. Motor activity was tested on the day of each morphine administration. After behavioral testing, brain tissue (striatum, hippocampus and prefrontal cortex) was collected for molecular experiments. The mRNA expression of orexin, dopamine and adenosine receptors was detected by qRT-PCR. In addition, the mRNA expression of markers such as GFAP and Iba-1 was analyzed by the same technique. SB-334867 inhibited the acquisition of morphine-induced motor activity sensitization in mice. During behavioral sensitization, the mRNA expression of orexin, dopamine and adenosine receptors as well as the expression of GFAP and Iba-1 were significantly altered, indicating that there is extensive interaction between orexin, dopamine, adenosine and glial cells in the mesolimbic system. In summary, the orexin system may be an effective means of inhibiting morphine-induced behavioral sensitization. [1]
To study the role of orexin-1 and orexin-2 receptor activity in ethanol self-administration, we used a high-drinking rodent model to evaluate compounds that differentially target orexin (OX) receptor subtypes in various self-administration paradigms. This study employed a two-bottle selection method to test the effects of the OX1 antagonist SB334867, the OX2 antagonist LSN2424100, and the mixed OX1/2 antagonist allomorexant (ACT-078573) on cage-borne ethanol intake in alcohol-loving (P) rats. In another experiment, the effects of SB334867, LSN2424100, and allomorexant on operant ethanol self-administration in P rats trained with a progressive ratio operant reinforcement program were assessed. In a third experiment, SB334867, LSN2424100, and allomorexant were administered to alcohol-loving C57BL/6J mice to investigate the effect of OX receptor blockade on ethanol intake in a binge-drinking (drinking in secret) model. In P rats subjected to long-term free choice of ethanol consumption, both SB334867 and allomorexant significantly reduced ethanol intake, but allomorexant also reduced water consumption, suggesting a non-specific effect on feeding behavior. In the progressive ratio operant conditioning experiment, LSN2424100 and almorexant reduced breakpoint and ethanol consumption in P rats, while the inactive enantiomer of almorexant and SB334867 had no significant effect on ethanol intake motivation. As expected, in the dark drinking model, mice injected with the vehicle exhibited a binge-drinking pattern. Compared with the control group injected with the vehicle, all three OX antagonists reduced ethanol intake and blood ethanol concentration, but SB334867 and LSN2424100 also reduced sucrose consumption in another group of mice, suggesting a non-specific effect. Overall, these results provide growing evidence that OX1 and OX2 receptor activity influences ethanol self-administration, although this effect may not be selectively targeted at ethanol intake. [2] The pharmacology of various peptide and non-peptide ligands in Chinese hamster ovary (CHO) cells stably expressing human orexin-1 (OX(1)) or orexin-2 (OX(2)) receptors was investigated by measuring intracellular calcium ([Ca(2+)](i)) using Fluo-3AM. Orexin A and orexin B increased intracellular calcium concentration [Ca(2+)](i) in CHO-OX(1) cells (pEC(50) 8.38±0.04 and 7.26±0.05, n=12, respectively) and CHO-OX(2) cells (pEC(50) 8.20±0.03 and 8.26±0.04, respectively, n=8, respectively). However, neuropeptide Y and secretin (10 pM - 10 μM) did not exhibit agonist or antagonist properties in either cell line. SB-334867-A (1-(2-methylbenzoxazol-6-yl)-3-[1,5]naphthidium-4-ylurea hydrochloride) inhibited orexin A (10 nM) and orexin B (100 nM) induced calcium responses (pK(B) 7.27±0.04 and 7.23±0.03, respectively, n=8), but had no effect on UTP (3 μM) induced calcium responses in CHO-OX(1) cells. SB-334867-A (10 μM) also inhibited OX(2) mediated calcium responses (32.7±1.9% inhibition compared to orexin A). SB-334867-A did not exhibit agonist properties in either cell line. In conclusion, SB-334867-A is a non-peptide OX(1) selective receptor antagonist. [3] |
| Molecular Formula |
C17H14NCLN5O2
|
|
|---|---|---|
| Molecular Weight |
319.32
|
|
| Exact Mass |
319.11
|
|
| Elemental Analysis |
C, 57.39; H, 3.97; Cl, 9.96; N, 19.68; O, 8.99
|
|
| CAS # |
249889-64-3
|
|
| Related CAS # |
SB-334867 free base; 792173-99-0
|
|
| PubChem CID |
6604926
|
|
| Appearance |
White to off-white solid powder
|
|
| Density |
1.472g/cm3
|
|
| Boiling Point |
450.5ºC at 760 mmHg
|
|
| Flash Point |
226.2ºC
|
|
| Vapour Pressure |
2.63E-08mmHg at 25°C
|
|
| Index of Refraction |
1.793
|
|
| LogP |
3.869
|
|
| tPSA |
92.94
|
|
| Hydrogen Bond Donor Count |
2
|
|
| Hydrogen Bond Acceptor Count |
5
|
|
| Rotatable Bond Count |
2
|
|
| Heavy Atom Count |
24
|
|
| Complexity |
462
|
|
| Defined Atom Stereocenter Count |
0
|
|
| SMILES |
O=C(NC1=CC(OC(C)=N2)=C2C=C1)NC3=CC=NC4=CC=CN=C43.Cl
|
|
| InChi Key |
AKMNUCBQGHFICM-UHFFFAOYSA-N
|
|
| InChi Code |
InChI=1S/C17H13N5O2.ClH/c1-10-20-12-5-4-11(9-15(12)24-10)21-17(23)22-14-6-8-18-13-3-2-7-19-16(13)14;/h2-9H,1H3,(H2,18,21,22,23);1H
|
|
| Chemical Name |
1-(2-methylbenzo[d]oxazol-6-yl)-3-(1,5-naphthyridin-4-yl)urea hydrochloride
|
|
| Synonyms |
|
|
| HS Tariff Code |
2934.99.9001
|
|
| Storage |
Powder -20°C 3 years 4°C 2 years In solvent -80°C 6 months -20°C 1 month Note: Please store this product in a sealed and protected environment, avoid exposure to moisture. |
|
| Shipping Condition |
Room temperature (This product is stable at ambient temperature for a few days during ordinary shipping and time spent in Customs)
|
| Solubility (In Vitro) |
DMSO: ~63 mg/mL (~197.3 mM)
Water: 1 mg/mL (Insoluble) Ethanol: ~3 mg/mL (~9.4 mM) |
|
|---|---|---|
| Solubility (In Vivo) |
|
| Preparing Stock Solutions | 1 mg | 5 mg | 10 mg | |
| 1 mM | 3.1317 mL | 15.6583 mL | 31.3165 mL | |
| 5 mM | 0.6263 mL | 3.1317 mL | 6.2633 mL | |
| 10 mM | 0.3132 mL | 1.5658 mL | 3.1317 mL |
*Note: Please select an appropriate solvent for the preparation of stock solution based on your experiment needs. For most products, DMSO can be used for preparing stock solutions (e.g. 5 mM, 10 mM, or 20 mM concentration); some products with high aqueous solubility may be dissolved in water directly. Solubility information is available at the above Solubility Data section. Once the stock solution is prepared, aliquot it to routine usage volumes and store at -20°C or -80°C. Avoid repeated freeze and thaw cycles.
Calculation results
Working concentration: mg/mL;
Method for preparing DMSO stock solution: mg drug pre-dissolved in μL DMSO (stock solution concentration mg/mL). Please contact us first if the concentration exceeds the DMSO solubility of the batch of drug.
Method for preparing in vivo formulation::Take μL DMSO stock solution, next add μL PEG300, mix and clarify, next addμL Tween 80, mix and clarify, next add μL ddH2O,mix and clarify.
(1) Please be sure that the solution is clear before the addition of next solvent. Dissolution methods like vortex, ultrasound or warming and heat may be used to aid dissolving.
(2) Be sure to add the solvent(s) in order.
|
|---|
 |
 |
Products are for research use only; We do not sell to patients
Copyright 2020 InvivoChem LLC | All Rights Reserved


 COA
